TCR-TRANSLATE
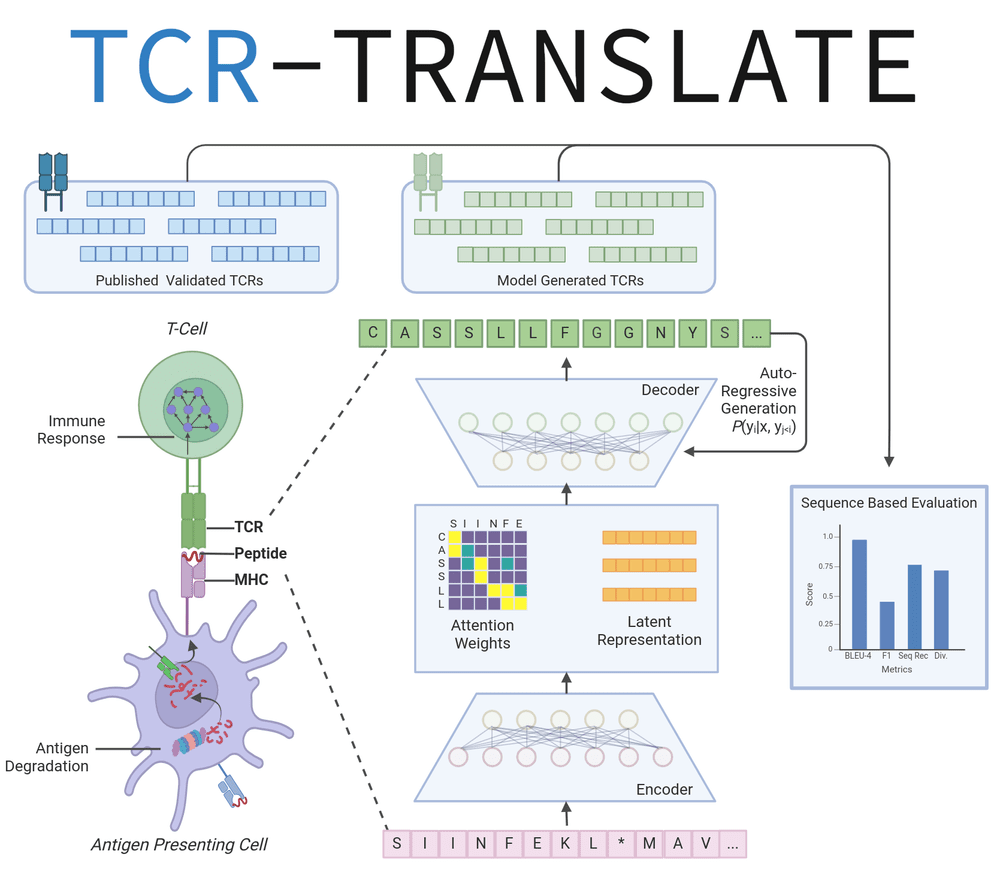
A sequence-to-sequence language model for antigen-specific TCR design that addresses data sparsity through techniques from low-resource machine translation.
PIRL uses and creates a variety of computational tools for our work in genomics and computational immunology. Many of these tools are carried over from previous work by the Vincent Lab at UNC, and OpenVax. Newer tools which are developed by PIRL will be listed below.
A sequence-to-sequence language model for antigen-specific TCR design that addresses data sparsity through techniques from low-resource machine translation.
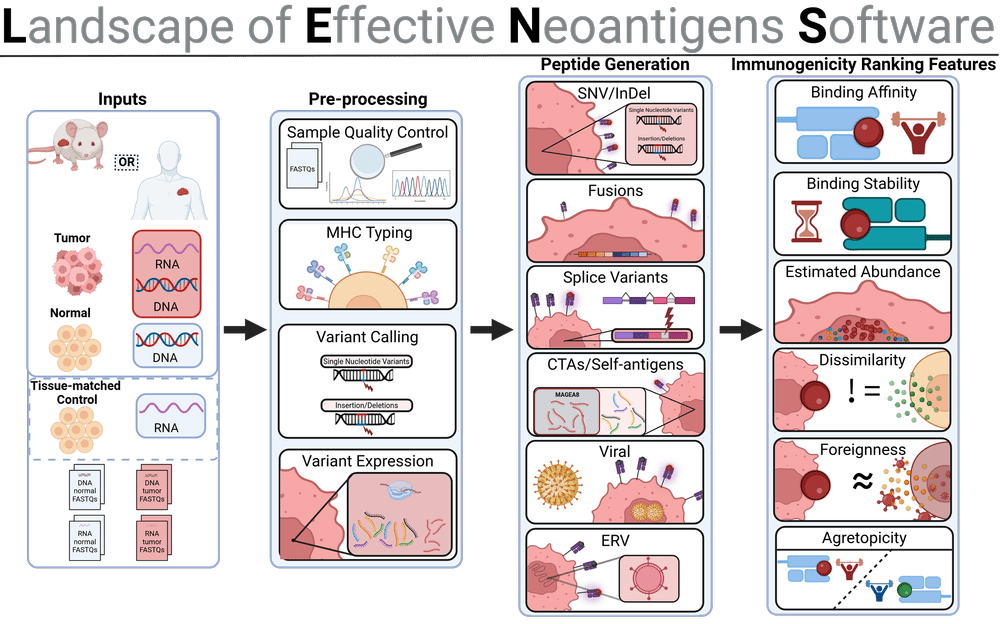
Tumor antigen prediction pipeline used to curate tumor-specific predicted MHC ligands from a variety of mutational and expression sources.
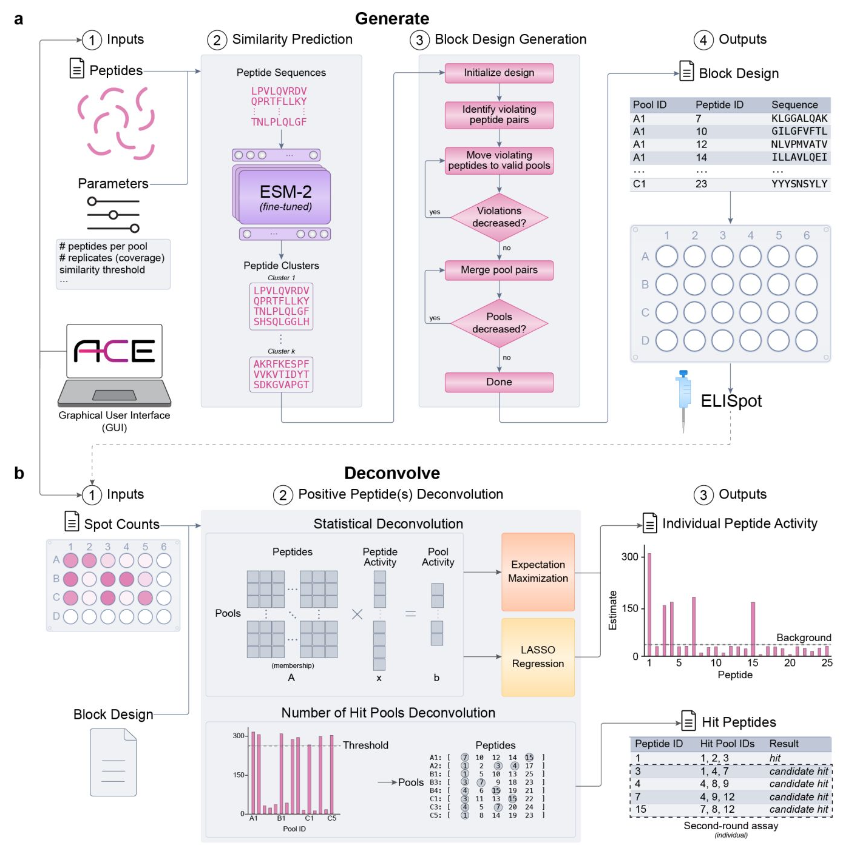
Multiplex ELISpot assay design and deconvolution tool for determining epitope-specific T-cell responses from a large number of candidate peptides.
A tricky part of constructing large immunological datasets is dealing with the chaotic variability in naming of MHC molecules across species and within different research communities. MHCgnomes is a parser for MHC nomenclature which uses several reference data sources and a robust parsing strategy to standardize MHC nomenclature from a variety of different naming schemes you might encounter in the wild.
Reproducible Analyses Framework and Tools: infrastructure for configuring and running reproducible Nextflow analyses.
A long-read toolkit for mutant proteoform prediction that integrates DNA and RNA variants, assembles transcripts, and translates them into annotated proteoforms and mutant peptides.
Pulled from github.com/pirl-unc when the site is built.
| Repository | Description | Language | Last push |
|---|---|---|---|
| weirdo | Metrics of immunological foreignness for candidate T-cell epitopes | Jupyter Notebook | |
| pirlygenes | Gene sets used for cancer immunotherapy expression analysis | Python | |
| presto | All of T-cell:pMHC prediction in one model | Python | |
| hitlist | Curated mass spectrometry evidence for MHC ligand data from IEDB and CEDAR | Python | |
| oncoref | Curated cancer reference data: ontology, TMB, incidence/mortality, and expression. Bottom-of-stack for the openvax/PIRL libraries. | Python | |
| pirl-unc.github.io | pirl.unc.edu | Python | |
| mhcgnomes | Parsing MHC nomenclature in the wild | Python | |
| mhcseqs | MHC sequences | Python | |
| io-resistance-genie | [WIP / experiment] Living literature review of resistance to immune checkpoint inhibitors | Python | |
| DoesExactoWorkYet | Simple end-to-end test of Exacto | Python | |
| vmwhere | exploring microsatellites in the human genome | Python | |
| tsarina | Tumor Specific Antigen discovery: personalized cancer immunotherapy targets from curated shared antigen data | Python | |
| trufflepig | Step-by-step RNA tumor analysis driven by pirlygenes gene sets — subsumes pirlygenes.analyze as many composable sub-commands with serializable intermediate state. | Python | |
| tcrsift | Python | ||
| exacto | Long-read toolkit for mutant proteoform prediction - somatic/germline DNA and RNA assembly, variant calling, integration, and translation | Rust | |
| singlecell_images | Docker files for images containing seurat + scCDC and seurat + decontx ( celda ) | Shell | |
| alcohol-immunotherapy | Data and code for "Alcohol use reduces the efficacy of anti-PD1 immunotherapy by disrupting anti-tumor immunity" | ||
| runplz | Tiny Modal-shaped job harness | Python | |
| raft-ui | RAFT web interface for generating manifests and configuration files. | HTML | |
| nexus | A command-line streamliner for Nextflow bioinformatics pipelines. | Nextflow | |
| aggrescan3d-slurm-workflow | Python | ||
| pdockq-slurm-workflow | Python | ||
| vstol | Variant Standardization, Tabulation, and Operations Library | Python | |
| io-shieldbreak | Python | ||
| squid-tui | Squid-TUI (Slurm QUeue Interactive Dashboard): Terminal UI (TUI) for organizing, filtering, and managing SLURM jobs. | Python | |
| golfy | Heuristic solver for pooled peptide experiment design | Jupyter Notebook | |
| perseo | Personalized cancer immunotherapy target selection from curated shared antigen data | Python | |
| hf-tcr | HuggingFace Scaffolding for EZ TCR Generation and Evaluation | Python | |
| TCRpMHCdataset | Python Class for Handling Many-to-Many Nature of TCR::pMHC Data | Python | |
| pirl-blog-notebooks | Jupyter notebooks accompanying PIRL Blog posts | Jupyter Notebook | |
| PRAME_in_NUT_Carcinoma | R | ||
| tahp | Tumor Antigen Heterogeneity Profiler | Python | |
| pdf-reports | Helping code for compiling plots and other outputs into presentation pdfs. | R | |
| ace | Deep learning-powered multiplexed ELISpot assay design and deconvolution of pool readouts for immunogenic peptide identification. | Jupyter Notebook | |
| datasetprep | Methods to make life easier when preparing datasets for analysis. | R | |
| lens_demo_workflow_outputs | HTML | ||
| binfotron | R | ||
| housekeeping | R | ||
| rserver-binfotron | Dockerfile | ||
| translate-pacbio-reads-for-bbn963 | Jupyter Notebook | ||
| longread-fusion-transcript-pipeline | R | ||
| lineardesign | docker image to run externally developed LinearDesign software | ||
| MiXCR | Dockerfile and script to run MiXCR | Dockerfile | |
| pvacitopes | Epitope selection from pVACtools output | Python | |
| bbn963_fusion_transcript_pipeline | A pipeline associated with the longread fusion pipeline used to extract fusions from our associated pacbio bbn963 data. | R | |
| NeoantigenProfiler | R | ||
| nutm1-fusions | Hunting for T-cell epitopes in recurrent NUTM1 somatic fusions | Jupyter Notebook | |
| lens | Landscape of Effective Neoantigens Software | ||
| yolotrim | Heuristic read trimming for PacBio IsoSeq data | Rust | |
| PostRNASeqAlign | R | ||
| nara | NUTM1 Associated Recurrent Antigens: possible T-cell targets on NUTM1 rearranged neoplasms | Jupyter Notebook | |
| rserver-immunesubtypes | hosts dockerfile for running immune subtypes package | Dockerfile | |
| variform | Variant isoform detection for large mutations in long-read RNA sequencing | Python | |
| python-project-template | Template of a basic Python project | Python | |
| coding-guidelines | Coding guidelines and recommendations for tools developed by PIRL | ||
| rserver | rserver | Shell | |
| PrePostMiXCR | Functions for writing commands and post processing the MiXCR pipeline. | R | |
| rserver-postproc-rnaseq | Dockerfile to make RStudio server for doing post processing of rnaseq data using the StarSalmon R-package | Dockerfile | |
| Landscape-and-Selection-of-Vaccine-Epitopes-in-SARS-CoV-2 | Code and data for "Landscape and Selection of Vaccine Epitopes in SARS-CoV-2" | Jupyter Notebook | |
| mHA-dockerfile | Dockerfile | ||
| lenstools | Dockerfile | ||
| command-line-tools | Tools helpful for working with data, nextflow, etc. | R | |
| picard | Adds magrittr to docker://broadinstitute/picard:2.27.5 to bypass error when using apptainer. | Dockerfile | |
| are-we-good | CI status of every project we develop and/or help maintain | ||
| HLAProfiler | R | ||
| StarSalmon | R | ||
| rserver-sc | RServer for single cell analyses. Contains monocle3, zinbwave and scRNAseq | Dockerfile | |
| VincentLabFormatter | R | ||
| stig | Synthetic T-cell Informatics Generator | Python | |
| fastmultiqc | Has environment and script for running fastqc and then multiqc on the results. | Shell | |
| trust_published | Dockerfile for running the original published version of TRUST. | Dockerfile | |
| trust | Dockerfile for running TRUST 3.0 | R | |
| MiXCR-2.1.7 | Dockerfile and script to run MiXCR 2.1.7 RNAseq | Java | |
| MiXCR-amplicon-2.1.7 | Dockerfile & script used for the diversity paper MiXCR amplicon runs. | Java | |
| Bowtie2 | Dockerfile and examples for using Bowtie2 | Shell | |
| tide | Dockerfile | ||
| rserver-impro | Dockerfile | ||
| making-sense-of-tesla | Jupyter Notebook | ||
| .github | Personalized Immunotherapy Research Lab at UNC | ||
| neoag | Mirror of vincentlaboratories / neoag | R | |
| fusion-transcript-workflow | |||
| ace-analysis | HTML | ||
| raft | Python | ||
| bbn963-mutations | Exploring long read WGS somatic variants for BBN963 cell line | Jupyter Notebook | |
| python-rust-template | The purpose of this repository is to test continuous integration using GitHub Actions | Python | |
| defense-against-the-dark-arts | Course website and resources for BCB 731 | HTML | |
| lbg-guide | Basic usage tips for the LBG cluster at UNC-CH | ||
| Rmbleven | An R utility for string comparisons using the mbleven algorithm written in Rcpp. | C++ | |
| tricking_nextflow_cache | This is a minimal code example demonstrating use of the clean_work_files.sh script to delete intermediate files in Nextflow workflows. | Nextflow | |
| blogs | Repositories for Code Associated with Blog Posts on PIRL Website | Jupyter Notebook | |
| immproper | T-cell epitope predictor | Python | |
| u87-ttf-mutational-signature | Mutational signature analysis of U87 glioma cells exposed to Tumor Treating Fields | Shell | |
| sv-web-parser-and-visualizer | Python | ||
| u87-u937-isoseq-demux | Demultiplexing of Iso-Seq cell line data | Python | |
| metaseq | Metadata format for collections of FASTA files containing genomic sequencing reads | ||
| SeqPipe | Reusable pipeline for sequence alignment and variant calling. | ||
| unc-sars-cov-2-sequencing | Analysis of SARS-CoV-2 viral sequencing data | Jupyter Notebook | |
| isovar-on-mouse-cell-lines-shell-scripts | Shell script that runs the RNA aligner star | Jupyter Notebook | |
| til-unc.github.io | Website for Translational Immunogenomics Lab | HTML | |
| klamp | SARS-CoV-2 detection from LamPORE sequencing data | Python | |
| cd8-tcell-epitope-prediction-data | Data for building predictive models of CD8+ T-cell epitopes | Jupyter Notebook |